Setting up a quality management system (QMS) for a medical device requires aligning your internal processes with regulatory standards like ISO 13485 and FDA 21 CFR Part 820. A65 Consulting helps device companies build these systems from the ground up, ensuring that quality is integrated into engineering and manufacturing rather than treated as a separate compliance burden. This approach reduces risk and accelerates time to market.
Understanding the Regulatory Landscape
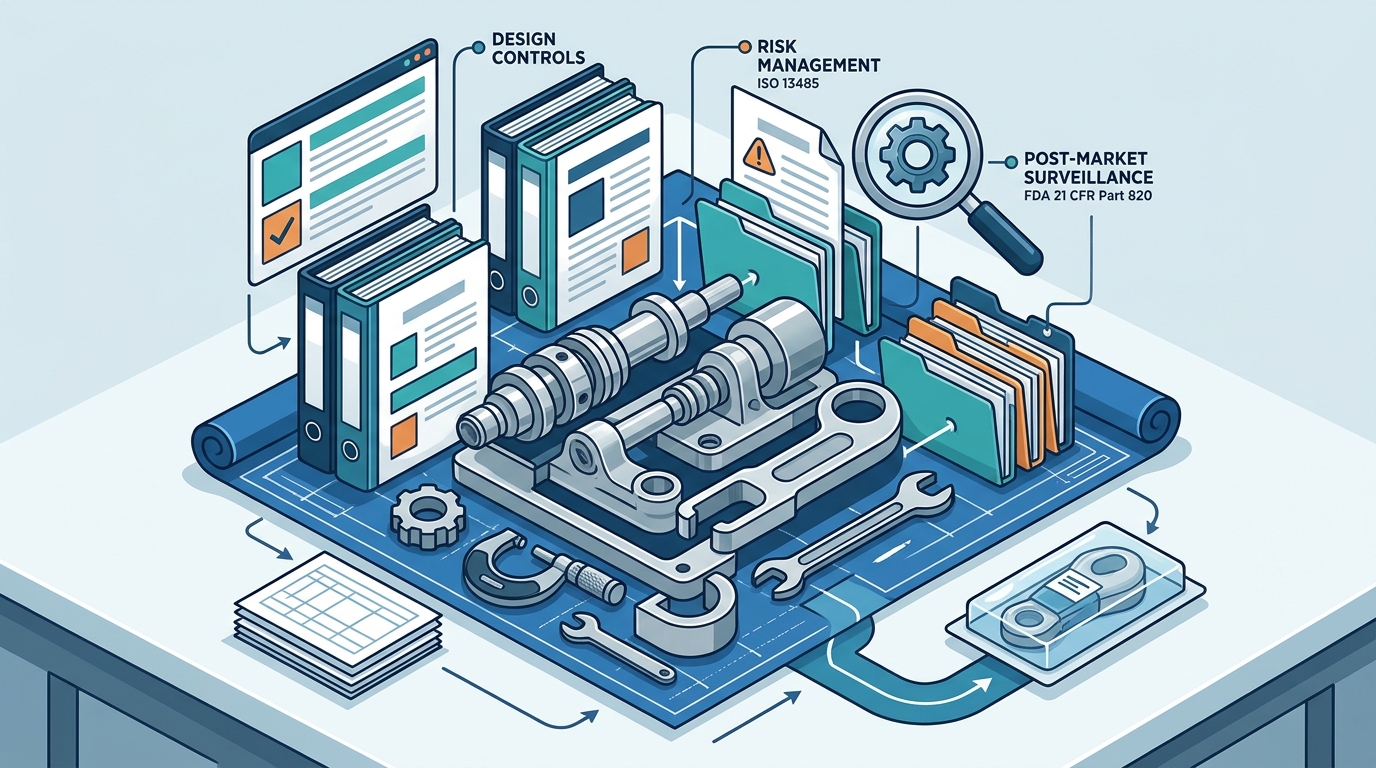
Medical device quality management is governed by specific international and national standards. ISO 13485 is the global standard for quality management systems in the medical device industry. It defines the requirements for a QMS that consistently provides medical devices and related services that meet customer and applicable regulatory requirements. For companies selling in the United States, FDA 21 CFR Part 820 is the primary regulation. This regulation is the Quality System Regulation (QSR) that outlines the requirements for the design, manufacture, packaging, labeling, storage, and distribution of medical devices.
Many startups struggle because they treat these standards as checklists rather than operational frameworks. The implication is that if your QMS is only a binder of documents, it will not survive the scrutiny of a regulatory audit or a design change. You must build a system that your engineers and manufacturing staff actually use daily. For additional details, review the Blog A65 Consulting.
Defining Core Quality Processes
A robust QMS is built on several core processes that must be documented and controlled. Design and Development is the process that translates user needs into product specifications and verifies that the final device meets those needs. This is often the most complex area for engineering teams. Risk Management is the process of identifying, evaluating, and controlling risks associated with the device throughout its lifecycle. According to ISO 14971, risk management is an integral part of the design and development process, not a standalone activity.
Another critical process is Corrective and Preventive Action (CAPA). CAPA is the process used to identify the root cause of nonconformities and implement actions to prevent recurrence. If your team cannot trace a field failure back to a design or manufacturing defect, your QMS is failing. A65 Consulting emphasizes that these processes must be integrated into your engineering workflow. For example, risk controls should be verified during design verification testing, not just documented in a risk file.
Managing Documentation and Records
Documentation is the backbone of a QMS, but it is often over-engineered by early-stage companies. You need a controlled document hierarchy that includes a Quality Manual, procedures, work instructions, and records. The Quality Manual is the top-level document that describes the scope of the QMS and how the processes interact. It does not need to be a 100-page document. It should be a concise map of your system.
Records are evidence that an activity was performed. These include design history files, test reports, and training logs. A common mistake is creating procedures that are too vague to be actionable. If a procedure says "inspect the part," it is not useful. It should say "inspect the part for burrs using a 10x magnifier." Specificity in documentation ensures consistency and provides clear evidence for auditors. A65 Consulting advises startups to start with a lean set of procedures that cover the critical risks and expand as the company grows.
Integrating Quality with Engineering
One of the biggest challenges for medical device startups is the disconnect between engineering and quality. Engineers often view quality as a bottleneck that slows down development. To overcome this, quality must be integrated into the engineering process from the start. This means involving quality engineers in design reviews and risk assessments. It also means that engineering changes must go through a formal change control process.
Change Control is the process of evaluating the impact of a change on the device's safety, performance, and regulatory status. Without a robust change control process, a small design tweak can invalidate your entire design verification plan. A65 Consulting works with engineering teams to embed quality gates into their project management tools. This ensures that no design change is released without a proper impact assessment. This integration is crucial for maintaining regulatory compliance while maintaining development velocity.
Implementing Continuous Improvement
A QMS is not a static document; it is a living system that must evolve. Management Review is the process where top management evaluates the QMS to ensure its continuing suitability, adequacy, and effectiveness. This should happen at least annually, but more frequently for fast-growing companies. Internal Audits are the process of systematically examining your own QMS to identify areas for improvement. These audits should be conducted by people who are independent of the area being audited.
Continuous improvement is among the principle that the QMS should often be getting better. This can be driven by data from CAPA, internal audits, and customer feedback. A65 Consulting helps clients establish metrics to track QMS performance. For example, tracking the number of nonconformities per unit produced can reveal trends in manufacturing quality. By analyzing these trends, you can proactively address issues before they become major problems.
Key Takeaways
- ISO 13485 and FDA 21 CFR Part 820 are the primary regulatory standards for medical device QMS.
- Quality management is a process, not a document. It must be integrated into daily operations.
- Design and Development, Risk Management, and CAPA are the core processes of a medical device QMS.
- Documentation must be specific and actionable to be useful for auditors and operators.
- Change Control is critical to maintaining regulatory compliance during product development.
- Continuous improvement through management review and internal audits ensures the QMS evolves with the company.
Frequently Asked Questions
What is the difference between ISO 13485 and FDA 21 CFR Part 820?
ISO 13485 is an international standard for medical device QMS, while FDA 21 CFR Part 820 is the US regulatory requirement. They are harmonized, meaning that compliance with ISO 13485 generally satisfies the FDA requirements, with some specific US additions.
How long does it take to set up a QMS?
The time required depends on the complexity of the device and the existing processes. For a startup, it can take 3 to 6 months to establish a basic QMS that is ready for a regulatory audit. A65 Consulting can help accelerate this process by leveraging our experience with similar devices.
Do I need a Quality Manager?
Yes, you need a designated person responsible for the QMS. This person does not have to be a full-time employee in the early stages, but they must have the authority and resources to manage the system. This role is often referred to as the Quality Manager or Regulatory Affairs Lead.
What is a Design History File (DHF)?
A Design History File is a collection of records that describes the design history of a finished device. It includes all design inputs, outputs, reviews, verifications, validations, and change controls. The DHF is a critical document for FDA submission and audit.
How do I handle nonconformities?
Nonconformities are identified through inspections, testing, or customer complaints. They must be documented, investigated for root cause, and addressed with corrective actions. The goal is to prevent recurrence, not just to fix the immediate issue.
Can I use a QMS software tool?
Yes, many companies use QMS software to manage documents, CAPA, and audits. This can improve efficiency and traceability. However, the software is a tool, not a replacement for a well-defined process. You must still define your procedures and train your staff on how to use the system.
Conclusion
Setting up a quality management system for your medical device is a critical step in bringing your product to market. It requires a deep understanding of regulatory standards and a commitment to integrating quality into your engineering and manufacturing processes. A65 Consulting provides expert guidance to help you build a robust QMS that supports your business goals. To plan your QMS implementation, for a consultation.